Expert-led consulting to achieve seamless approvals
across global markets.
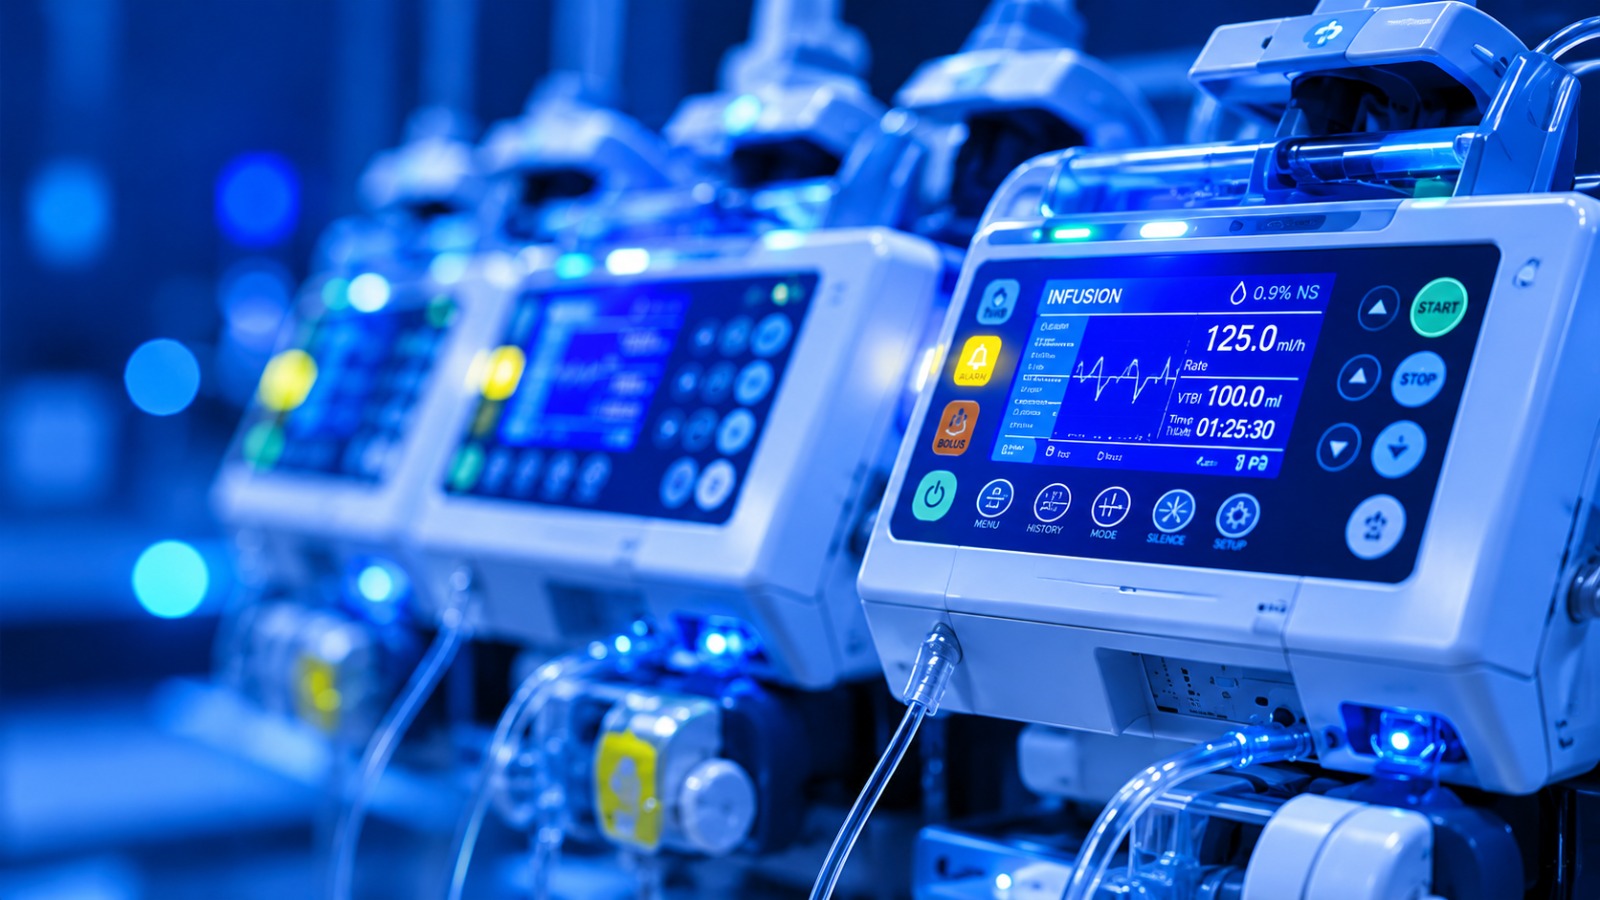
Global Regulatory Expertise
CE, USFDA, CDSCO & more
End-to-End Support
From strategy to approval
Expert Consultants
Industry veterans by your side
Global Market Access
Helping you reach more markets

Tacit MedTek is a customer-focused consulting firm delivering regulatory
and quality compliance solutions for healthcare product manufacturers worldwide.
We offer end-to-end support—from startups to multinational companies—covering
regulatory strategy, system implementation, training, licensing, and certifications.
Trusted by 500+ Healthcare Innovators Worldwide


Our Speciality
ISO certification is an external credential validating that a business meets globally recognized standards for quality, safety, or efficiency.
Custom MDR solutions designed to accelerate approvals and market success.
Maven Profcon handles the most efficient and comprehensive Regulatory solutions for Registrations of Cosmetics, Renewals, Amendments, Label Reviews Etc.
AI-SaMD (Artificial Intelligence Software as a Medical Device) represents software designed to perform medical functions without being hardware-dependent.
End-to-end CE marking support under MDR & IVDR for EU market access.
Comprehensive cyber security measures for medical devices and digital health solutions.
Information security and privacy are closely related but distinct concepts.
Pharmaceutical services encompass the full spectrum of activities required to discover, develop, manufacture, and distribute medications.
Biologicals services encompass the discovery, development, manufacturing, and regulatory compliance of products derived from living organisms, such as vaccines, gene therapies, and monoclonal antibodies.
Cosmetic services include a wide array of surgical and non-surgical procedures designed to enhance physical appearance and boost self-esteem.
Nutraceutical services span the lifecycle of dietary supplements and functional foods, bridging the gap between food and medicine.
Our Gallery
FAQ
Of course. To make your compliance journey easier, Tacit Medtek provides end-to-end assistance with documentation, regulatory submissions, and communication with CDSCO and other regulatory bodies.
Yes. The Medical Devices Rules of 2017 mandate that, depending on the device's risk class, all medical devices must have a license before being manufactured or imported.
No, ISO 13485:2016 is not required to obtain a manufacturing license under MDR-2017; nonetheless, it is highly advised because it complies with regulatory requirements.
Yes, it is not required to indicate a device's expiration date or shelf life if it is given in a non-sterile state.
Technical paperwork, clinical evaluation, conformance assessment, and adherence to EU MDR/IVDR are all part of CE marking. Startups can get help from Tacit Medtek during the whole CE certification procedure.
It is necessary for devices that interact with the human body directly or indirectly. ISO 10993 requirements must be followed when conducting tests, and they must be revalidated if materials or procedures change.
Yes, if the materials, processes, and suppliers are unchanged, and supported with justification and a biological risk assessment.
Yes. If you are a foreign manufacturer, you are required to designate an Indian Authorized Agent to manage communication and regulatory submissions.
No, a separate application, supporting documentation, and costs are needed for each actual manufacturing site.
If your company's constitution changes, you must notify the licensing authority and reapply for a license within the allotted time (usually 180 days).
A medical device that has undergone processing to eradicate all microbiological life and is packaged
to
remain sterile until it is opened or damaged is considered sterile. The device's sterility must be
explicitly stated on the label, along with the sterilizing technique that was
employed.
However, a
non-sterile medical device is one that is provided without being sterilized. The label of the devices
must
explicitly say that it is "non-sterile" and provide instructions for correctly sterilizing it before
use
if that is the plan.
No, Chapter XI of MDR-2017 exempts Class A non-sterile and non-measuring medical devices from wholesale licensing.
In addition to clearly stating that the equipment is non-sterile, the package must reduce microbiological contamination and contain instructions for sterilizing appropriate for the planned approach prior to patient use.
Yes. Any changes must be reported to the licensing authority within 45 days, and the licensee has 180 days from the date of the constitution change to submit an application under MDR-2017. In these cases, unless a new license is granted or the application is rejected by the licensing authority, the current license will be considered valid.
Yes. Within 45 days of the constitution change, the licensee has 180 days to file an application under MDR-2017, and any modifications must be communicated to the licensing authority. In these situations, the existing license will be deemed valid unless a new license is issued or the licensing body rejects the application.
In order to indicate that a medical device is sold freely in the countries of Australia, Canada, Japan, the European Union, the United Kingdom, or the United States of America, the Free Sale Certificate (FSC) granted to the legal manufacturer must include brand names (registered under the Trade Marks Act, 1999) for the device in Form MD-14. If the brand name is not listed in the FSC, it should be listed on Form MD-14 as "Not Applicable."
Contact Us
Blogs


Top 10 Mistakes Startups Make in Medical Device Regulatory Compliance
Medical device startups frequently fail by treating regulatory compliance as a late-stage box-ticking exercise rather than an integral part of development. Key mistakes include neglecting early quality management systems (QMS), confusing validation with verification, and underestimating documentation needs, often leading to costly delays, rejected submissions, and failed commercialization.

US FDA 510(k) vs CE Marking vs CDSCO: Key Differences Explained
The US FDA 510(k), CE Marking (EU MDR), and CDSCO (India) pathways differ primarily in their focus on substantial equivalence, clinical evidence, and speed to market. 510(k) focuses on equivalence to a US predicate, CE Marking emphasizes technical compliance and safety, and CDSCO focuses on safety and regulatory compliance within India.
Connect With Us